Hydrogen Atom Wavefunctions: Radial Distribution, Nodes and Shapes
A rigorous conceptual guide to Schrödinger solutions, nodal surfaces, radial probability density, and orbital geometries for graduate chemistry aspirants.
From the Schrödinger Equation to Quantum Numbers and Wavefunctions
The hydrogen atom occupies a foundational place in quantum mechanics because it is the only atomic system for which the non-relativistic time-independent Schrödinger equation can be solved analytically in closed exact form. For higher-tier competitive examinations such as IIT-JAM, CSIR-NET, GATE, and CUET-PG, questions frequently test whether an aspirant understands the mathematical origins of quantum numbers rather than merely memorizing formulas.
The hydrogenic system consists of a positively charged nucleus with atomic number Z and a single electron of mass me moving under a central Coulombic potential:
V(r) = -Z e² / (4πε₀ r)
Because the potential depends strictly on the radial distance r between the nucleus and the electron, Cartesian coordinates (x, y, z) are computationally unwieldy. Transforming the Hamiltonian into spherical polar coordinates (r, θ, φ)—where r is radial distance, θ is the polar (colatitudinal) angle ranging from 0 to π, and φ is the azimuthal angle ranging from 0 to 2π—allows separation of variables.
The total wavefunction, ψ(r, θ, φ), factorizes into two independent functions:
ψn,l,m(r, θ, φ) = Rn,l(r) · Yl,m(θ, φ) = Rn,l(r) · Θl,m(θ) · Φm(φ)
Here, the wavefunctions [तरंग फलन] are governed by three spatial quantum numbers arising directly from the boundary conditions required to keep ψ single-valued, continuous, and normalizable:
- Principal Quantum Number (n): Takes values n = 1, 2, 3, ... It determines the quantized electronic energy in one-electron systems (En = -Z²RH / n²) and dictates the radial extent or physical size of the electron cloud.
- Azimuthal or Orbital Angular Momentum Quantum Number (l): Takes integer values l = 0, 1, 2, ..., (n - 1). It quantizes the total orbital angular momentum magnitude, given by |L| = √[l(l + 1)] ħ, and determines orbital shape.
- Magnetic Quantum Number (m or ml): Takes (2l + 1) integer values from -l to +l. It describes the spatial orientation of the angular momentum vector along a chosen quantization axis (typically the z-axis), where Lz = m ħ.
Radial vs Angular Wavefunctions: Mathematical Structure and Physical Meaning
Understanding the anatomy of the radial wavefunction Rn,l(r) is essential for solving graph-interpretation and root-finding questions in national examinations. The general mathematical solution for the radial component of hydrogen-like atoms is expressed through associated Laguerre polynomials:
Rn,l(r) = - Nn,l · (2Zr / na₀)l · e-Zr / na₀ · Ln+l2l+1(2Zr / na₀)
where a₀ is the Bohr radius (approximately 0.529 Å) and Nn,l is a normalization constant. Notice three critical structural elements in every radial wavefunction:
- The Power Term (rl): The term rl governs behavior as r → 0. For s-orbitals (l = 0), r⁰ = 1, which means R(0) has a finite, non-zero maximum value at the nucleus. For any orbital with l > 0 (such as p, d, f), rl drives the wavefunction strictly to zero at the nucleus: R(0) = 0. This physical barrier arises from the centrifugal potential term l(l + 1)ħ² / (2μr²), which repels the electron from the nuclear origin when angular momentum is non-zero.
- The Polynomial Term: A polynomial of degree (n - l - 1) in terms of r. The real, positive roots of this polynomial represent radial nodes—distances where the radial wavefunction crosses zero.
- The Exponential Decay Term (e-Zr / na₀): This damping term ensures the wavefunction decays smoothly toward zero as r → ∞, satisfying the condition of normalizability (square-integrability).
Conversely, the angular wavefunction Yl,m(θ, φ) consists of associated Legendre polynomials in cos(θ) combined with the azimuthal factor eimφ. Because eimφ yields complex functions for m ≠ 0, chemists construct real-valued atomic orbitals (such as px, py, dxy, dx²-y²) by taking normalized linear combinations of the degenerate positive and negative m states.
Nodes in Wavefunctions: Radial Nodes, Angular Nodes, and Nodal Surfaces
A node [नोड] is a point, plane, or surface where the electronic wavefunction passes through zero (ψ = 0) and changes its algebraic sign (+ to - or vice versa). Consequently, the probability density [प्रायिकता घनत्व] at a nodal surface is strictly zero. In three dimensions, nodes are categorized into two mutually exclusive types:
1. Angular Nodes
Angular nodes arise when the angular spherical harmonic equals zero: Yl,m(θ, φ) = 0. Because the angular function depends entirely on l, the total count of angular nodes is always equal to l. Angular nodes manifest in two geometric forms:
- Nodal Planes: Flat planes passing directly through the nucleus where the electron probability is zero. For example, a pz orbital has its nodal plane at z = 0 (the xy-plane). A dxy orbital has two nodal planes: x = 0 (the yz-plane) and y = 0 (the xz-plane).
- Nodal Cones: Conical surfaces passing through the nucleus. The dz² orbital has l = 2, so it must possess 2 angular nodes. However, it does not possess two planar nodes. Instead, setting its angular factor (3cos²θ - 1) = 0 yields cos(θ) = ±1/√3, which corresponds to two symmetric conical surfaces oriented at approximately 54.74° and 125.26° relative to the z-axis.
2. Radial Nodes
Radial nodes are concentric spherical shells centered at the nucleus where Rn,l(r) = 0 for 0 < r < ∞. The number of radial nodes is governed by the expression:
Radial Nodes = n - l - 1
For instance, solving the radial factor for a 2s orbital (where R2s is proportional to (2 - Zr/a₀)) gives a single root at r = 2a₀/Z. For a 3s orbital, the quadratic polynomial yields two distinct positive roots, creating two concentric nodal spheres.
Total Nodal Topology
Summing both types gives the total number of nodes in any orbital:
Total Nodes = (n - l - 1) + l = n - 1
The table below summarizes the distribution of nodes across representative atomic orbitals up to the fourth shell:
| Orbital | Principal (n) | Azimuthal (l) | Radial Nodes (n - l - 1) | Angular Nodes (l) | Total Nodes (n - 1) | Geometric Nature of Nodes |
|---|---|---|---|---|---|---|
| 1s | 1 | 0 | 0 | 0 | 0 | No nodes |
| 2s | 2 | 0 | 1 | 0 | 1 | 1 spherical shell (r = 2a₀/Z) |
| 2p | 2 | 1 | 0 | 1 | 1 | 1 planar surface passing through nucleus |
| 3s | 3 | 0 | 2 | 0 | 2 | 2 concentric spherical shells |
| 3p | 3 | 1 | 1 | 1 | 2 | 1 spherical shell + 1 planar surface |
| 3d | 3 | 2 | 0 | 2 | 2 | 2 planes (or 2 conical surfaces for dz²) |
| 4s | 4 | 0 | 3 | 0 | 3 | 3 concentric spherical shells |
| 4p | 4 | 1 | 2 | 1 | 3 | 2 spherical shells + 1 planar surface |
| 4d | 4 | 2 | 1 | 2 | 3 | 1 spherical shell + 2 planes/cones |
| 4f | 4 | 3 | 0 | 3 | 3 | 3 angular surfaces (planar or conical) |
Aspirants studying through our curated study material should practice deducing these nodal counts directly from the polynomial factors of wavefunctions rather than relying solely on memory.
Radial Probability Density vs Radial Distribution Function
One of the most persistent confusions among graduate students is the distinction between radial probability density and the radial distribution function (RDF) [त्रिज्यीय वितरण फलन]. Competitive exam questions exploit this conceptual boundary systematically.
Radial Probability Density: [R(r)]²
The square of the radial wavefunction, [Rn,l(r)]², represents the probability of finding the electron per unit volume at a precise mathematical point located at distance r from the nucleus. For an s-orbital (l = 0):
- [R1s(0)]² is at its absolute maximum at r = 0 (the nuclear center).
- As r increases, [R1s(r)]² decays monotonically and exponentially toward zero.
The Radial Distribution Function: 4πr²[R(r)]²
In physical chemistry, we rarely want to know the probability at a single microscopic point. Instead, we ask: What is the probability of finding the electron inside a thin spherical shell of thickness dr at a distance r from the nucleus, integrated over all angles (0 ≤ θ ≤ π, 0 ≤ φ ≤ 2π)?
The differential volume element in spherical polar coordinates is:
dV = r² sin(θ) dr dθ dφ
Integrating over the complete angular domain gives:
∫0π sin(θ) dθ ∫02π dφ = 2 · 2π = 4π
Therefore, the volume of a thin spherical shell at distance r with thickness dr is 4πr² dr. Multiplying this spherical shell volume by the probability density gives the Radial Distribution Function, denoted as P(r) or D(r):
P(r) dr = 4πr² [Rn,l(r)]² dr
Why Does RDF Drop to Zero at the Nucleus?
Consider the 1s electron. At r = 0, the probability density [R(r)]² is maximum. However, the spherical shell factor 4πr² evaluates to 4π(0)² = 0. Physically, a sphere of radius zero has zero volume. Consequently, the probability of locating the electron inside a shell of zero thickness at the nucleus is strictly zero:
P(0) = 4π(0)² · [R(0)]² = 0
As r moves away from zero, 4πr² increases quadratically, while [R(r)]² decays exponentially. The product of an increasing function and a decreasing exponential produces a curve that starts at zero, rises to a distinct maximum, and then decays back asymptotically to zero.
Peak Topology and the Penetration Effect
The total number of peaks (maxima) in a radial distribution curve is governed by the relation:
Number of RDF Peaks = n - l
Of these (n - l) peaks, exactly (n - l - 1) are minor inner peaks, and 1 is the principal outer peak. Compare the 2s and 2p orbitals in hydrogen-like systems:
- 2s Orbital (n=2, l=0): Has 2 - 0 = 2 peaks. It displays a small inner peak close to the nucleus, a radial node at r = 2a₀/Z, and a large primary peak farther out.
- 2p Orbital (n=2, l=1): Has 2 - 1 = 1 peak. It has no radial nodes and rises to a single maximum.
Although the principal maximum of 2p lies closer to the nucleus than the outer maximum of 2s, the small inner peak of the 2s electron penetrates much closer to the nucleus than the 2p distribution ever reaches. This quantum mechanical penetration effect directly explains why s-electrons experience a higher effective nuclear charge (Zeff) and lower energy than p-electrons in multi-electron atoms.
Most Probable Radius (rmp) vs Average Radius (⟨r⟩)
Numerical and theoretical problems in CSIR-NET and GATE regularly test the mathematical distinction between the most probable distance and the expectation (mean) distance.
Derivation of Most Probable Radius for 1s Orbital
The most probable radius, rmp, represents the distance at which the radial distribution function P(r) attains its maximum value. To find it, take the first derivative of P(r) with respect to r and set it to zero:
P(r) = 4πr² · [2(Z/a₀)3/2 e-Zr/a₀]² = 16π (Z/a₀)³ · r² e-2Zr/a₀
Differentiating with respect to r using the product rule:
dP/dr = 16π (Z/a₀)³ [ 2r e-2Zr/a₀ - (2Z/a₀) r² e-2Zr/a₀ ] = 0
Factoring out common terms:
2r e-2Zr/a₀ [ 1 - (Zr / a₀) ] = 0
Excluding trivial boundaries (r = 0, where P is minimal, and r → ∞), the non-trivial root is:
rmp = a₀ / Z
For a neutral hydrogen atom (Z = 1), rmp = a₀ = 0.529 Å. The most probable distance predicted by Schrödinger wave mechanics matches Bohr's first orbit radius exactly. For any orbital where l = n - 1 (such as 1s, 2p, 3d, 4f), the general formula for the most probable radius is:
rmp = n² a₀ / Z
The Expectation Value: Average Radius ⟨r⟩
The average or mean distance ⟨r⟩ is the quantum mechanical expectation value calculated using the operator r̂ over all space:
⟨r⟩ = ∫ ψ* r ψ dV = ∫0∞ r · P(r) dr = ∫0∞ 4πr³ [R(r)]² dr
Evaluating the standard integral for the 1s orbital of a hydrogen-like atom yields:
⟨r⟩1s = (3/2) · (a₀ / Z) = 1.5 a₀ / Z
For general quantum states (n, l), the expectation value is given by the general expression:
⟨r⟩n,l = (a₀ / 2Z) · [ 3n² - l(l + 1) ]
For n = 2 states in hydrogen (Z = 1):
- For 2s (l = 0): ⟨r⟩2s = (a₀ / 2) [3(4) - 0] = 6 a₀
- For 2p (l = 1): ⟨r⟩2p = (a₀ / 2) [3(4) - 2] = 5 a₀
This mathematical proof confirms that on average, a 2s electron resides further from the nucleus than a 2p electron, even though its inner radial peak allows it to penetrate closer.
Orbital Shapes and Boundary Surfaces
Atomic orbitals cannot be enclosed by rigid physical walls. Because wavefunctions decay exponentially, an orbital extends infinitely into space. To represent orbitals pictorially, physical chemists employ boundary surface diagrams (enclosing a defined probability volume, typically 90% or 95%):
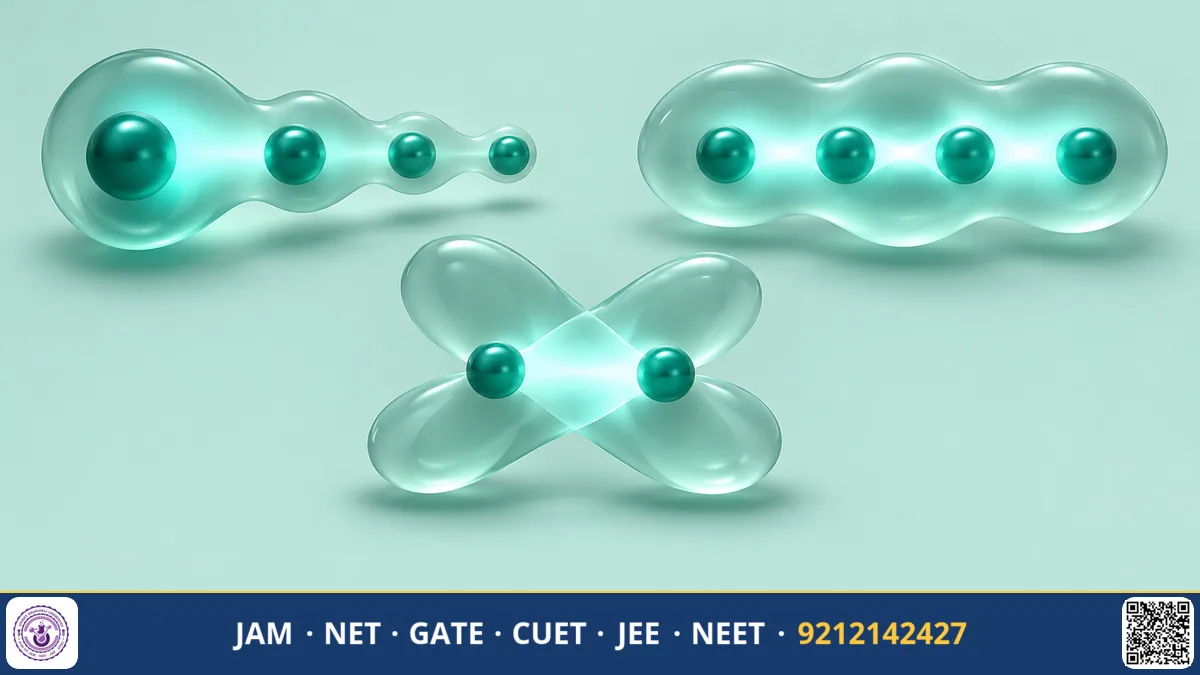
- s-Orbitals (l = 0, m = 0): The angular wavefunction Y0,0 = 1/√(4π) is a constant. Because there is no dependence on angles θ or φ, all s-orbitals are spherically symmetric. As n increases, the outer boundary surface expands, enclosing concentric spherical nodes.
- p-Orbitals (l = 1, m = 0, ±1): Dumb-bell shaped with two lobes separated by a nodal plane passing through the nucleus. The lobes carry opposite algebraic signs of the wavefunction (+ and -). These signs represent mathematical wave phase, not electrostatic charge. Phase matching is fundamental to constructive and destructive interference in molecular orbital theory.
- d-Orbitals (l = 2, m = 0, ±1, ±2): Four of the five d-orbitals (dxy, dyz, dxz, dx²-y²) exhibit cloverleaf geometries with four alternating lobes and two perpendicular nodal planes. The fifth, dz², has two vertical lobes along the z-axis surrounded by a toroidal doughnut-shaped ring in the xy-plane, accompanied by two conical nodal surfaces.
Typical Exam Questions and Problem-Solving Strategies for Competitive Exams
In national competitive examinations such as CSIR-NET, IIT-JAM, GATE, and CUET-PG, atomic structure and quantum mechanics form core areas of the physical chemistry syllabus. Exam question formats vary: IIT-JAM features Multiple Choice Questions (MCQs), Multiple Select Questions (MSQs), and Numerical Answer Type (NAT) questions; GATE relies on MCQs and NATs; and CSIR-NET includes high-weightage conceptual problems in Part C.
Rather than relying on arbitrary third-party claims regarding fixed marks or topic recurrence ratings, serious aspirants should evaluate syllabus-based areas that are commonly examined across official examination cycles. We recommend verifying the current pattern and section weightages directly from the official notification brochures issued by the organizing institutes. Aspirants should build their own tally by solving official previous year question papers and practicing with standard topic-wise test papers.
Standard Problem Archetypes
- Identifying Orbitals from Polynomials: You are provided a radial expression, such as R(r) = C · r² · (6 - Zr/a₀) · e-Zr/3a₀. Inspect the power of r at the front: here it is r², so l = 2 (a d-orbital). Inspect the polynomial: it is linear in r, so it has 1 radial node. Using n - l - 1 = 1 with l = 2 gives n - 2 - 1 = 1, which implies n = 4. The orbital is 4d.
- Calculating Node Coordinates: To find the radial node of R2s, set the polynomial factor (2 - Zr/a₀) = 0, yielding r = 2a₀/Z. For 3s, set the quadratic factor (27 - 18σ + 2σ²) = 0 (where σ = Zr/a₀) and solve using the standard quadratic formula.
- Evaluating Ratio of Maximum Densities: Comparing rmp values across hydrogenic ions (H, He⁺, Li²⁺, Be³⁺) using rmp = a₀/Z. For Li²⁺ (Z = 3), rmp = a₀/3.
- Graph Matching: Distinguishing plots of ψ vs r, ψ² vs r, and 4πr²ψ² vs r. Always check two boundary conditions: behavior at r = 0 (only s-orbitals have non-zero ψ and ψ²; all RDFs start at zero) and the number of times the curve touches or crosses the horizontal axis.
For systematic guidance on quantum chemistry and physical chemistry problem solving, students can contact our academic desk at 9212142427 to explore our classroom-centre and online classes.
Frequently Asked Questions
Why is the radial distribution function zero at the nucleus (r = 0) even for an s-orbital where the wavefunction is non-zero?
The radial distribution function is given by P(r) = 4πr²[R(r)]². While the radial wavefunction R(0) and probability density [R(0)]² have non-zero maximum values at the nucleus for s-orbitals, the geometric shell factor 4πr² evaluates strictly to zero when r = 0. Physically, the volume of a spherical shell of radius zero is zero, meaning there is zero physical volume at a singular point to contain probability.
What is the mathematical difference between the most probable radius and the average radius of a 1s electron?
The most probable radius (rmp) is the distance where the radial distribution function achieves its peak value (mode), found by setting d[P(r)]/dr = 0, which gives rmp = a₀/Z (0.529 Å for hydrogen). The average radius ⟨r⟩ is the quantum expectation value (mean) calculated as ∫ r P(r) dr, which yields 1.5 a₀/Z. The average radius is larger than the most probable radius because the distribution curve has a long exponential tail extending outward, shifting the mean away from the mode.
How do you determine the number of angular nodes for a dz² orbital?
The number of angular nodes depends solely on the azimuthal quantum number l. For any d-orbital, l = 2, so the dz² orbital has exactly 2 angular nodes. However, unlike the other four d-orbitals which have planar nodes, the angular factor of dz² is proportional to (3cos²θ - 1). Setting this factor to zero yields cos(θ) = ±1/√3 (approximately 54.74° and 125.26°), which defines two concentric conical nodal surfaces meeting at the nucleus.
Why do s-orbitals have non-zero probability density at the nucleus while p and d orbitals have zero?
The radial wavefunction near the origin behaves proportionally to rl. For s-orbitals, l = 0, so r⁰ = 1, resulting in a finite, non-zero value at r = 0. For p (l = 1), d (l = 2), and f (l = 3) orbitals, the rl term drives the wavefunction to zero at r = 0. Physically, any state with non-zero angular momentum experiences a centrifugal repulsive potential that prevents the electron from existing at the nuclear origin.
How many peaks are observed in the radial distribution curve of a 3p orbital?
The total number of peaks in any radial distribution function is given by n - l. For a 3p orbital, n = 3 and l = 1, so the number of peaks is 3 - 1 = 2. These consist of one minor inner peak and one major principal outer peak, separated by exactly one radial node (n - l - 1 = 3 - 1 - 1 = 1).
Are atomic orbital wavefunctions degenerate in one-electron systems?
In one-electron hydrogen-like atoms (such as H, He⁺, Li²⁺), the electronic energy depends strictly on the principal quantum number n (En = -Z²RH / n²) when relativistic fine-structure effects are neglected. Therefore, all subshells within the same principal shell (for example, 2s and 2p, or 3s, 3p, and 3d) are completely degenerate. In multi-electron atoms, electron-electron repulsions and differential penetration effects lift this subshell degeneracy, splitting energies such that Es < Ep < Ed < Ef.
Preparing for a chemistry entrance exam?
ABC Chemistry runs focused IIT-JAM, CSIR-NET, GATE and CUET-PG Chemistry coaching through live online classes for students across India.
Call / WhatsApp: 9212142427